Cyclic Voltammetry in Electrochemistry
Published:
Updated October 10, 2026.
There was always a cyclic voltammetry (CV) scan running on the lab computer (remotely controlled by my colleagues) every time I passed by. I got really curious about the cyclic voltammogram, especially why there were two $y$ (current) values corresponding to one $x$ (potential) value. What made me more curious was that my colleagues could show each other the voltammogram and talk science about it. I was too shy to ask how they did it (don’t be like me), so I decided to figure it out myself, not only because I was curious but also because, if I wanted to perform photoelectrochemistry experiments on plasmonic nanoparticles someday, I would need to know how CV works and what it can tell me. So I am writing this tutorial to educate myself.
Preliminary Knowledge
Chemical Equilibrium
For a simple chemical reaction
\[\mathrm{A}\rightleftarrows \mathrm{B}\]The equilibrium constant is defined as $K=\frac{[\mathrm{B}]}{[\mathrm{A}]}$, and once the equilibrium is reached the concentrations $[\mathrm{A}]$ and $[\mathrm{B}]$ do not change with time.
Kinetics
Equilibrium, however, does not mean that the reaction is “frozen”; it means that the forward reaction rate $v_f$ equals the backward reaction rate $v_b$ so the concentration of reactants and products does not change over time. Assuming the forward and backward reactions are both first-order reactions, then the reaction rates and the equilibrium condition can be written as:
\[v_f =k_f[\mathrm{A}],\qquad v_b=k_b[\mathrm{B}],\] \[v_f=v_b\Rightarrow k_f[\mathrm{A}]=k_b[\mathrm{B}],\]where $k_f$ and $k_b$ are the rate constants of forward and backward reactions, respectively. Both the equilibrium constant (a thermodynamic concept) and the kinetics are important as the former tells us what equilibrium looks like and the latter tells us how fast we can get there.
Activation Energy and the Arrhenius Equation
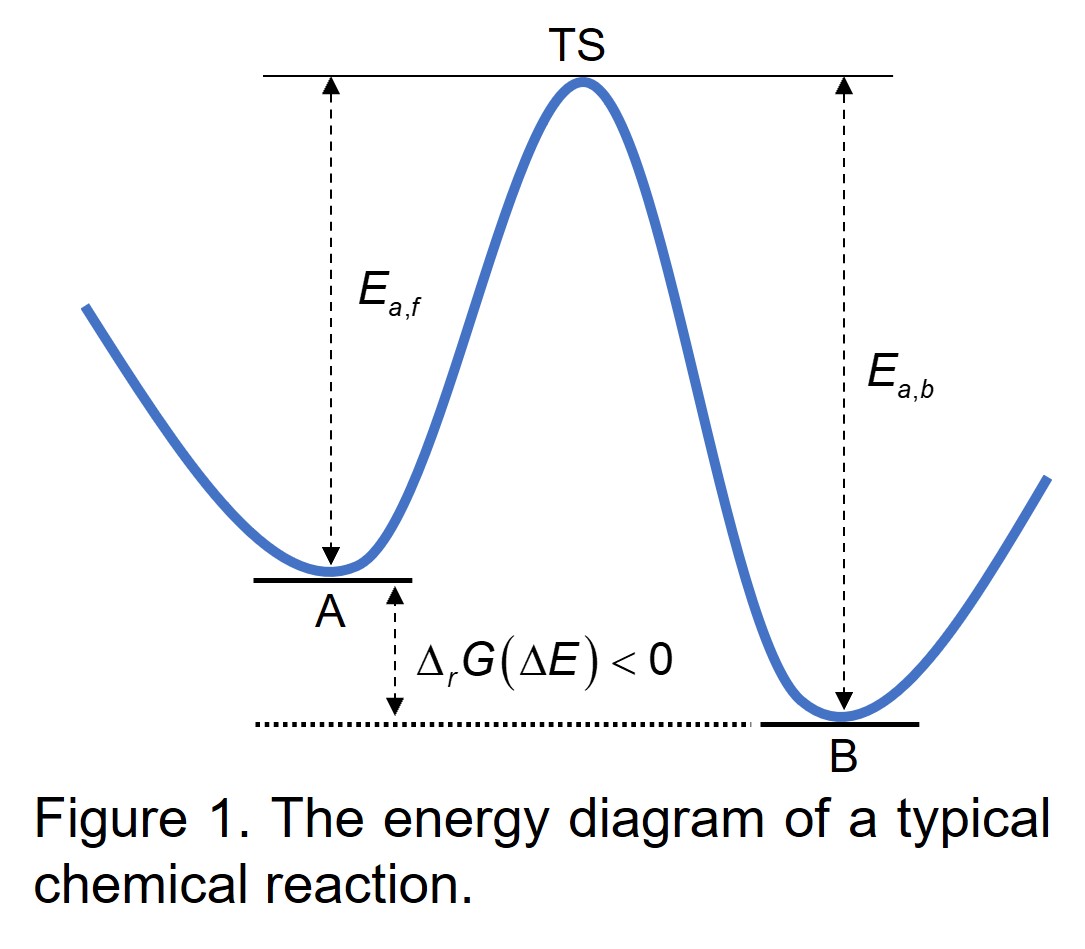
In most cases, there is a “hill” that reactant A needs to surpass before forming product B even if the reaction is energetically favorable $(\Delta_rG<0)$. The peak of the “hill” is usually represented by a transition state (TS) or activated complex, and we can therefore define activation energies for forward $(E_{a,f})$ and backward $(E_{a,b})$ reactions. This is the basis of the transition state theory (Figure 1).
The Arrhenius equation states that the rate constant is related to the activation energy and absolute temperature through
\[k=Ae^{\frac{-E_a}{RT}}\]where $A$ is a pre-exponential factor; it contains everything that we don’t know (lol) about the rate constant and ideally it should be temperature-insensitive. One can imagine that the rate constant would be greater (the reaction is faster) if the activation energy is smaller (or the temperature is higher), provided other reaction parameters are kept the same.
The Nernst Equation
The potential of a redox $(\mathrm{Ox}/\mathrm{Red})$ couple is characterized by the Nernst equation, which links the potential to the concentration of $\mathrm{Ox}$ and $\mathrm{Red}$ species. The Nernst equation in the general case is:
\[\mathrm{Ox}+n\mathrm{e^-}\rightleftarrows \mathrm{Red},\] \[E=E^0+\frac{RT}{nF}\ln\frac{C_\mathrm{Ox}}{C_\mathrm{Red}},\]where $C_\mathrm{Ox}$ and $C_\mathrm{Red}$ are the concentrations of $\mathrm{Ox}$ and $\mathrm{Red}$ species. The potential evaluated in the Nernst equation is also a thermodynamic property, but it turns out to also affect the reaction rates through the transition state theory (Figure 1): the ratio between $\mathrm{Ox}$ and $\mathrm{Red}$ species alters the relative position of $\mathrm{Ox}$ (A) and $\mathrm{Red}$ (B) on the potential energy surface (PES); therefore, it also alters the activation energies felt by $\mathrm{Ox}$ and $\mathrm{Red}$ (will be explained in detail below).
The above preliminary knowledge should put us on the same page and can be found in many undergraduate chemistry textbooks. Now, we start to derive the cyclic voltammogram.1–3
Electron Transfer Process (A General Description)
Since the redox reactions involve electron transfer, it wouldn’t hurt if we first study the electron transfer process in general (Figure 2). The simplest scenario is the one-electron transfer process that reduces $\mathrm{Ox_1}$ to its reduced form $\mathrm{Red_1}$.
\[\mathrm{Ox_1}+\mathrm{e^-}\rightarrow \mathrm{Red_1}\]Imagine we are electrons: why do we sometimes flow from A to B and sometimes from B to A? What is the driving force behind that? Well, simply put, we always want to flow to somewhere we feel more comfortable (low potential energy). This is universal regardless of whether it is a homogeneous electron transfer or heterogeneous electron transfer (Figure 2): To reduce $\mathrm{Ox_1}$ to $\mathrm{Red_1}$, the electrons may be injected from another reducing agent $\mathrm{Red_2}$ where the electrons are more uncomfortable (high potential energy) or from an electrode that is purposely maintained at a negative potential.
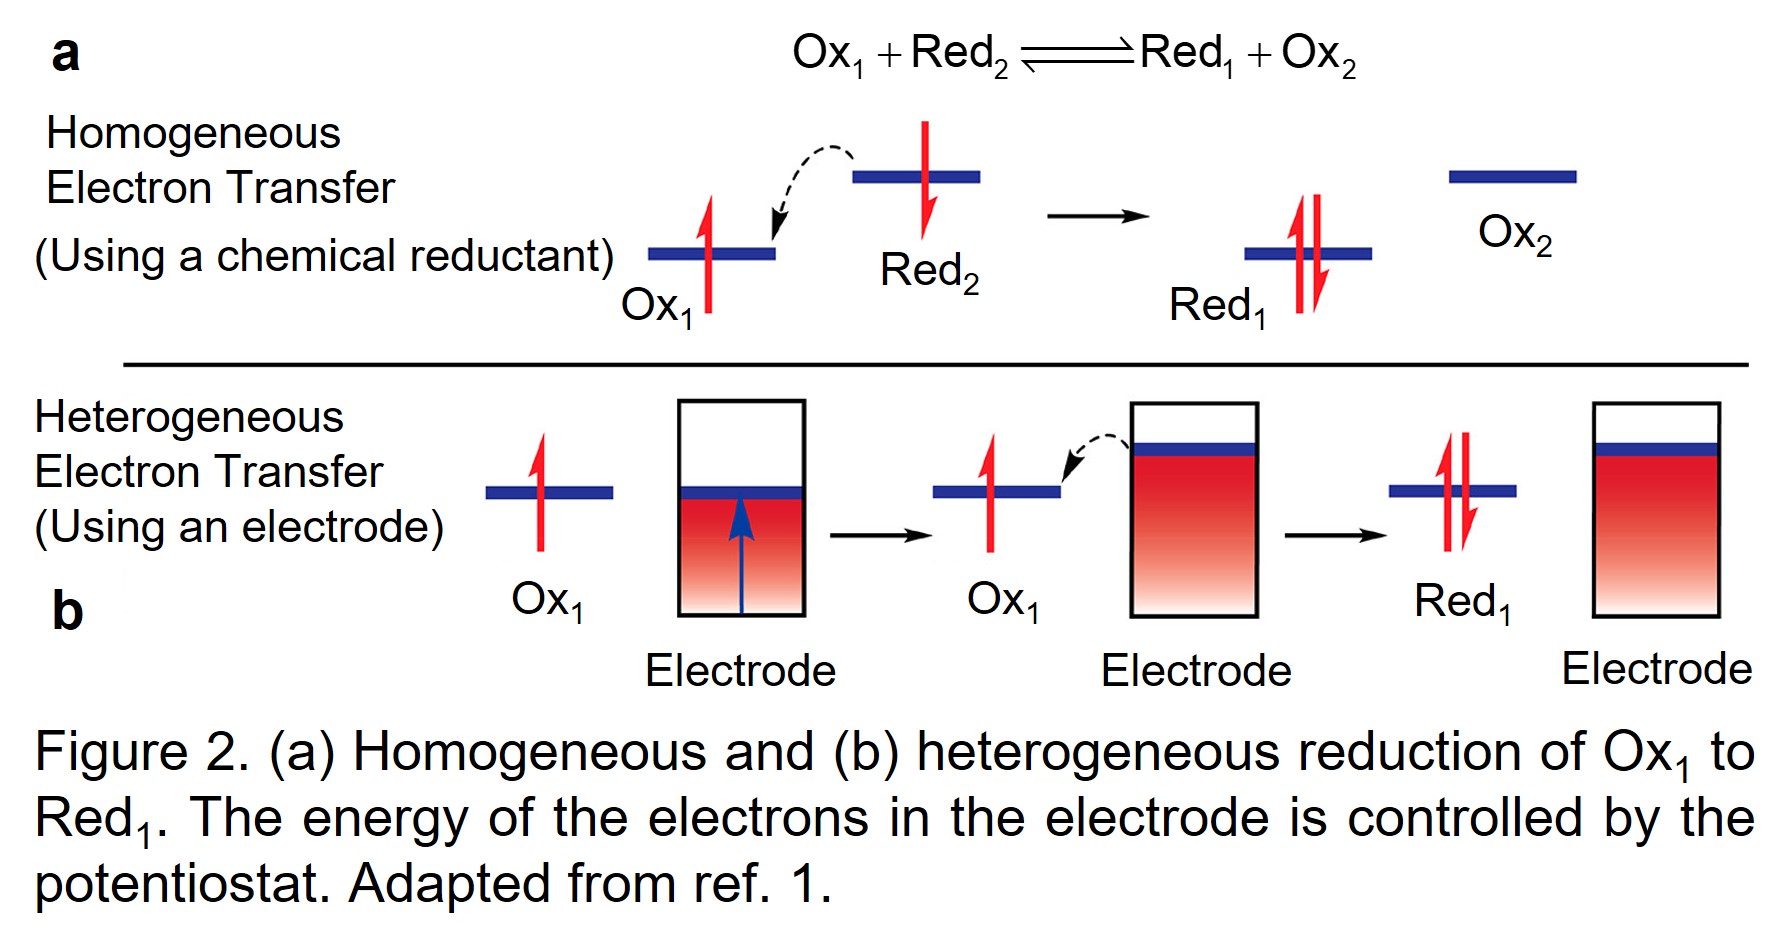
At this stage, it is important for us to keep this in mind: 1) electrons tend to flow from high energy to low energy, 2) for electrodes, positive potential tends to lower electron energy (electrons are more comfortable), thus preventing electrons from flowing out of the electrode, and vice versa.
Diffusion in an Electrochemical Cell
Now we are ready to see what my colleagues usually do in the lab: constructing an electrochemical (EC) cell and performing analysis such as CV. A representative EC cell (3-electrode configuration) is shown in Figure 3. As the name suggests, it contains three electrodes: the working electrode is the one we are primarily interested in as it drives the targeted redox reactions (of either the electrolyte or the electrode itself), the counter electrode is just for circuit connection purposes (it can extract or inject electrons into the electrolyte solution easily), and the reference electrode tells us what the potential of the working electrode is. That is why, when reporting a potential, we usually write vs. reference electrode.
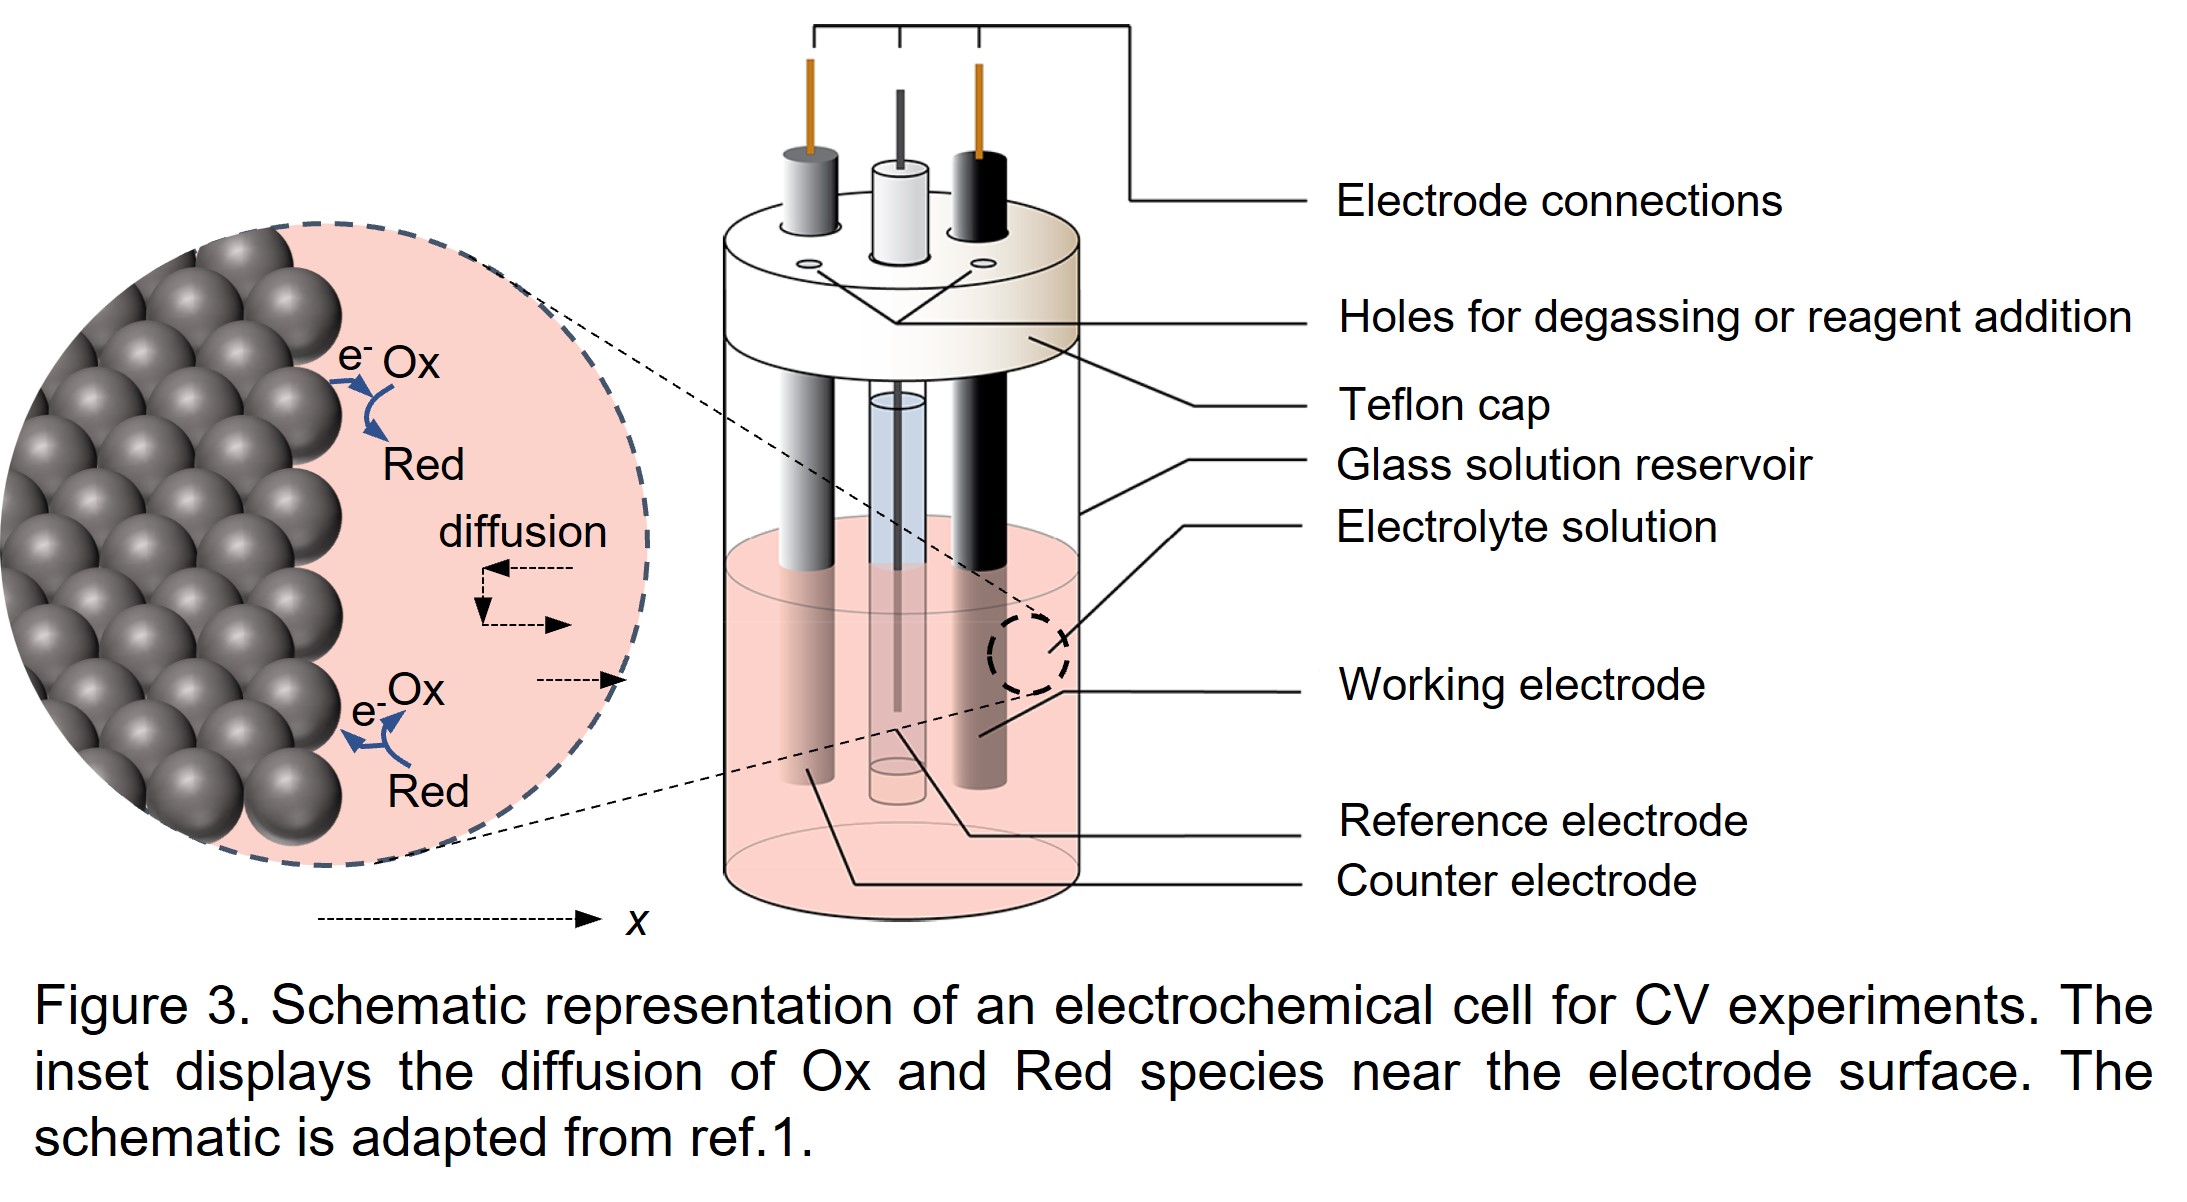
There is one key difference between an EC cell and an ordinary electric circuit: the current measured in an EC cell is dictated by the reaction rates of oxidation of $\mathrm{Red}$ (injecting electrons into the electrode) and reduction of $\mathrm{Ox}$ (extracting electrons from the electrode) occurring at the electrode surface, while the current measured in an ordinary electric circuit is determined by the voltage drop and the resistance (Ohm’s law). In an EC cell, therefore, the currents depend on the reaction rates, which are further related to the concentrations of $\mathrm{Ox}$ and $\mathrm{Red}$ species and the potential provided by the working electrode (see below). What’s important is that besides the reactions that consume or produce redox couples, both $\mathrm{Ox}$ and $\mathrm{Red}$ are subject to diffusion in the EC cell (see inset in Figure 3).
If, for simplicity, we treat the diffusion in one dimension only, that is, we assume both $\mathrm{Ox}$ and $\mathrm{Red}$ only diffuse perpendicular to the electrode surface and the electrode surface is perfectly smooth, then we can use Fick’s second law of diffusion (eq. 8) to describe the diffusion of $\mathrm{Ox}$ and $\mathrm{Red}$ away from and toward the surface.
\[\frac{\partial C}{\partial t}=D\frac{\partial^2 C}{\partial x^2},\]where $D$ is the diffusion coefficient. The direction of diffusion is from high concentration to low concentration.
Applied Potential Modifies Activation Energy
There is one significant question remaining, that is, how does the applied electric potential alter the PES (or equivalently, the energy profile along the reaction coordinate)? To illustrate this, we can consider a reduction process in which the reactant is $\mathrm{Ox}+\mathrm{e}^-$ (assume it is a one-electron, one-step process) and the electrons are from the working electrode. As we saw, a positive potential tends to lower electron energy while a negative potential raises it, so we can plot three scenarios in Figure 4, wherein the applied electric potential is referenced to the standard (or formal) potential $E^0$ of the redox couple under consideration.
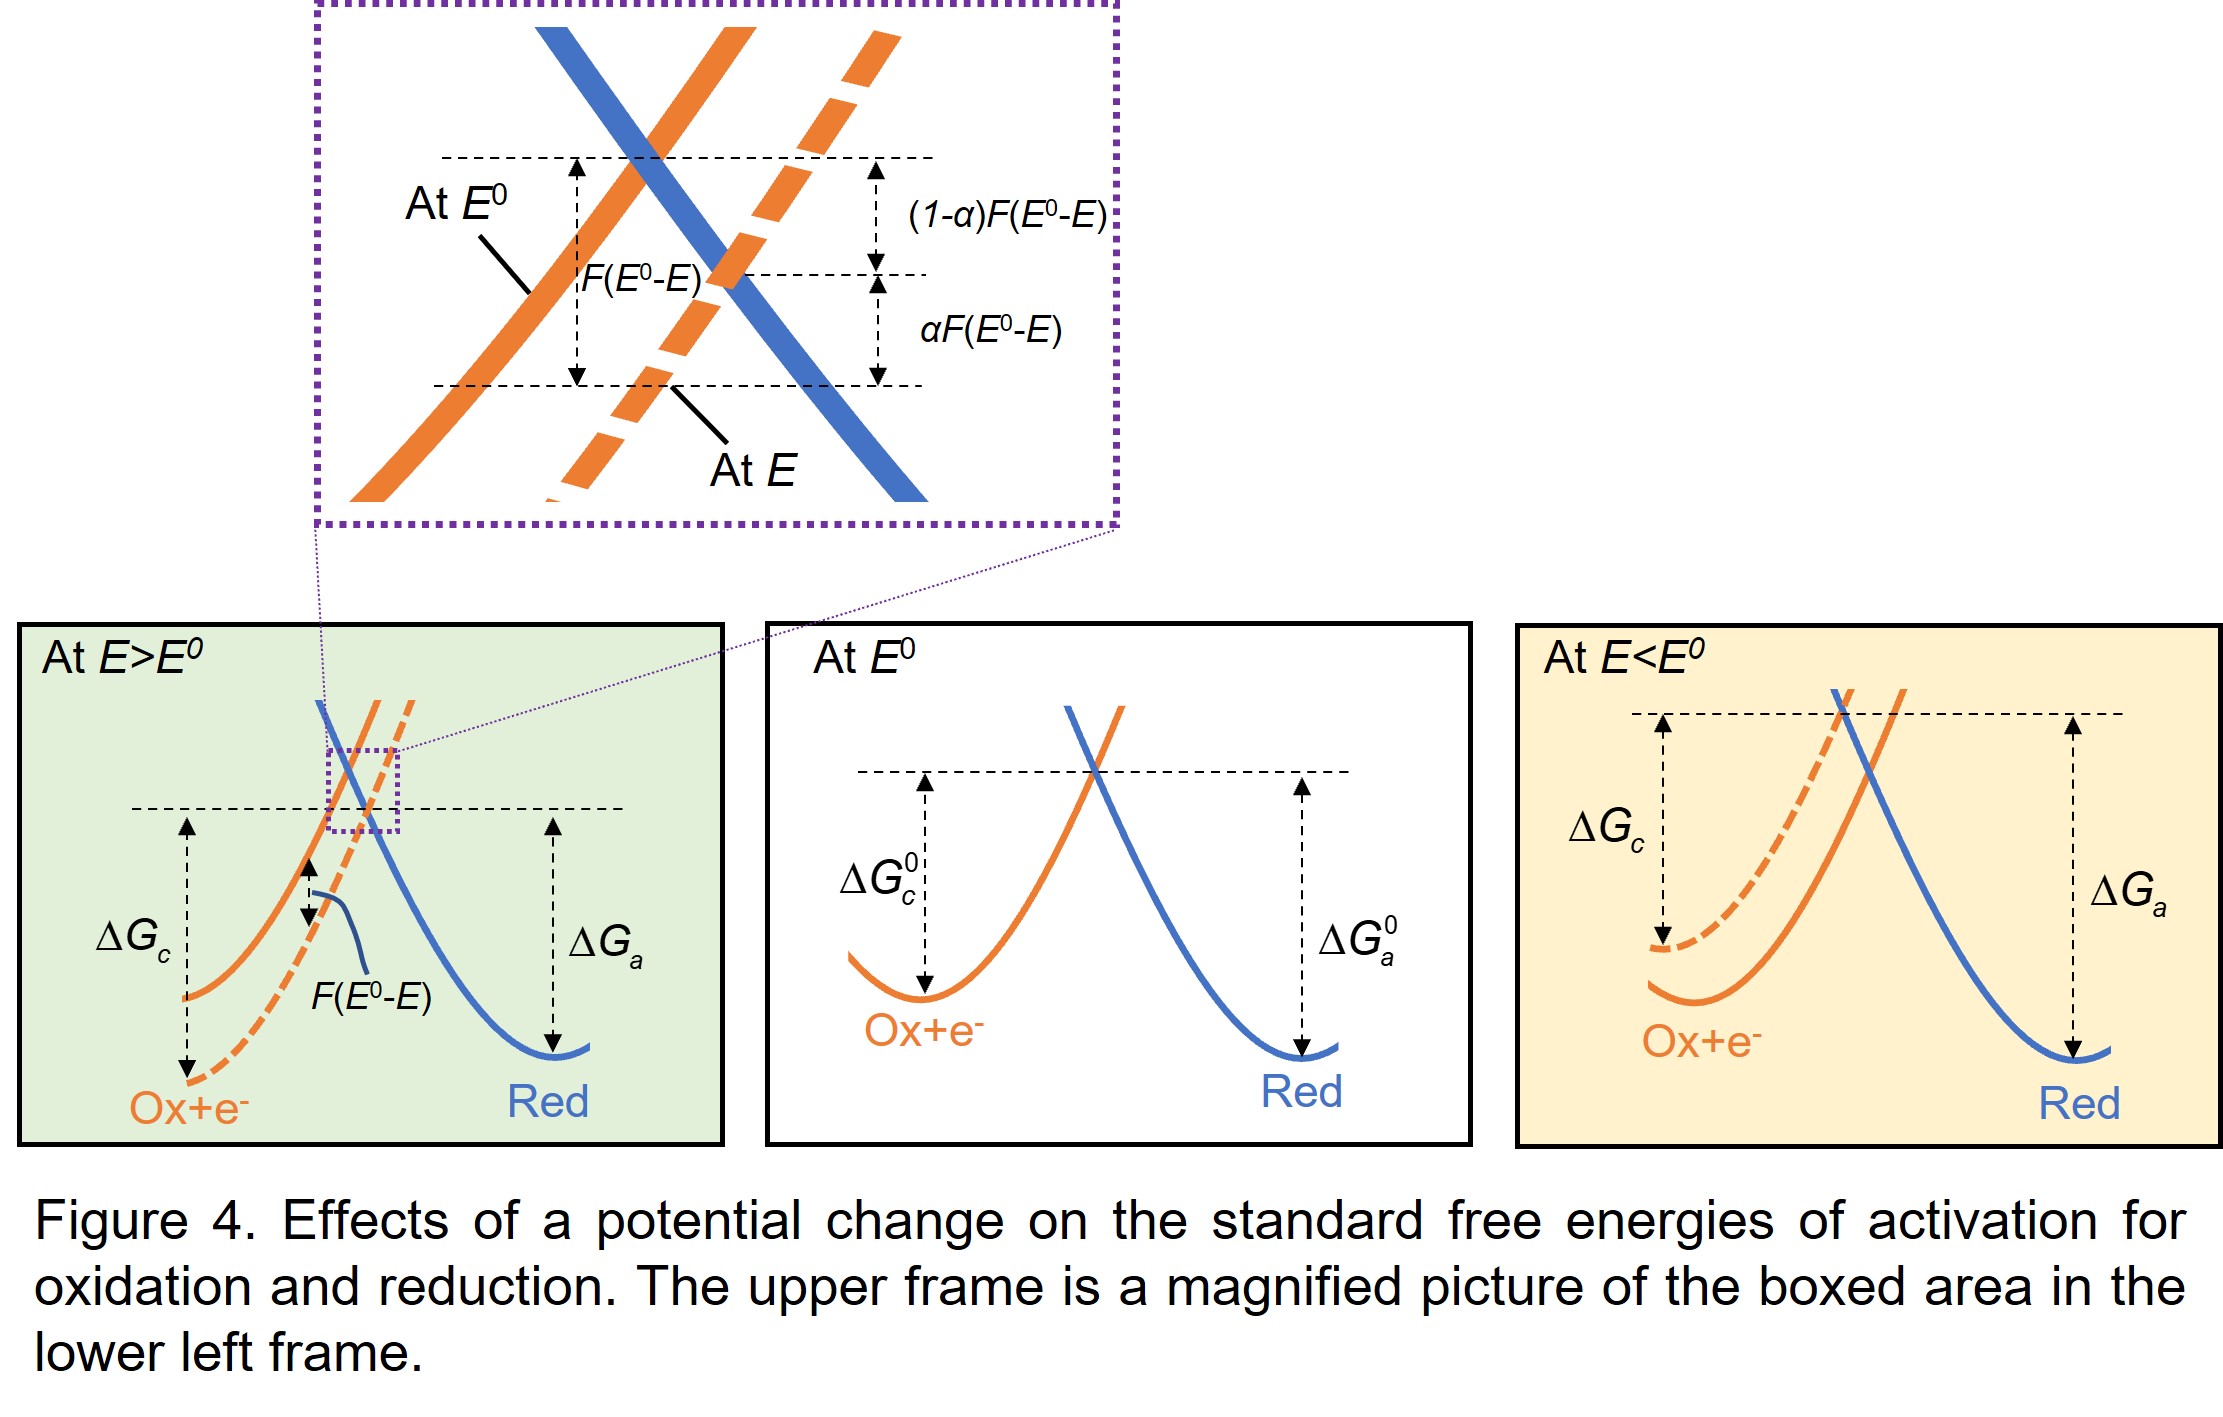
To avoid confusion, I have replaced activation energy $E_a$ with the free energy of activation $\Delta G$ (they are the same thing here), and the subscripts $c$ and $a$ stand for cathode (where reduction happens) and anode (where oxidation happens), respectively. The lower-left and middle panels of Figure 4 show that if the potential is changed by $\Delta E>0$ to a new value, the relative energy of the electron resident on the electrode changes by $-F\Delta E=-F(E-E^0)$; hence the $\mathrm{Ox}+\mathrm{e^-}$ curve moves down by that amount. Apparently (I hate this word), the barrier for reduction, $\Delta G_c$, has become greater than $\Delta G_c^0$, and the barrier for oxidation, $\Delta G_a$, has become less than $\Delta G_a^0$ by a fraction of the total energy change. Let us call that fraction $1-\alpha$, where $\alpha$, the transfer coefficient, can range from zero to unity.2
Therefore, we have the following two expressions for the activation energies for anodic and cathodic barriers:
\[\Delta G_a=\Delta G_a^0-(1-\alpha)F(E-E^0),\] \[\Delta G_c=\Delta G_c^0+\alpha F(E-E^0),\]If we assume the rate constants $k_f$ and $k_b$ have the Arrhenius form, we have
\[k_f=A_fe^{-\Delta G_c/RT}=A_fe^{-\Delta G_c^0/RT}e^{-\alpha F(E-E^0)/RT}=k_f^0e^{-\alpha F(E-E^0)/RT}=k^0e^{-\alpha F(E-E^0)/RT},\] \[k_b=A_be^{-\Delta G_a/RT}=A_be^{-\Delta G_a^0/RT}e^{(1-\alpha) F(E-E^0)/RT}=k_b^0e^{(1-\alpha) F(E-E^0)/RT}=k^0e^{(1-\alpha) F(E-E^0)/RT},\]The last $^”=^”$ in the above two equations is simply because $E^0$ can be chosen at the formal potential of the redox pair of interest, which gives $k_f^0=k_b^0=k^0$.
Final Step: The Point Method
Now we have everything we need to know to evaluate the current (the $y$ values) in a CV measurement. However, evaluating the diffusion using eq. 8 turns out to be difficult, so we use the point method3 to evaluate diffusion in the discrete form:
\[\frac{C_j-C_i}{\Delta t}=\frac{D[C_{i-1}-2C_i+C_{i+1}]}{\Delta x^2}\] \[C_j=C_i+\lambda(C_{i-1}-2C_i+C_{i+1})\]where $\lambda=D\Delta t/\Delta x^2$, and the distance (measured from the electrode surface) and time increments, $\Delta x$ and $\Delta t$, come from the point method (Figure 5a). In Figure 5b, we display the potential sweep in one cycle during CV measurement: the working electrode’s potential is swept from a high potential to a low potential (or vice versa), and then back to its original value. The CV measurement gets its name since during the potential sweep, the current due to redox reactions is measured simultaneously.
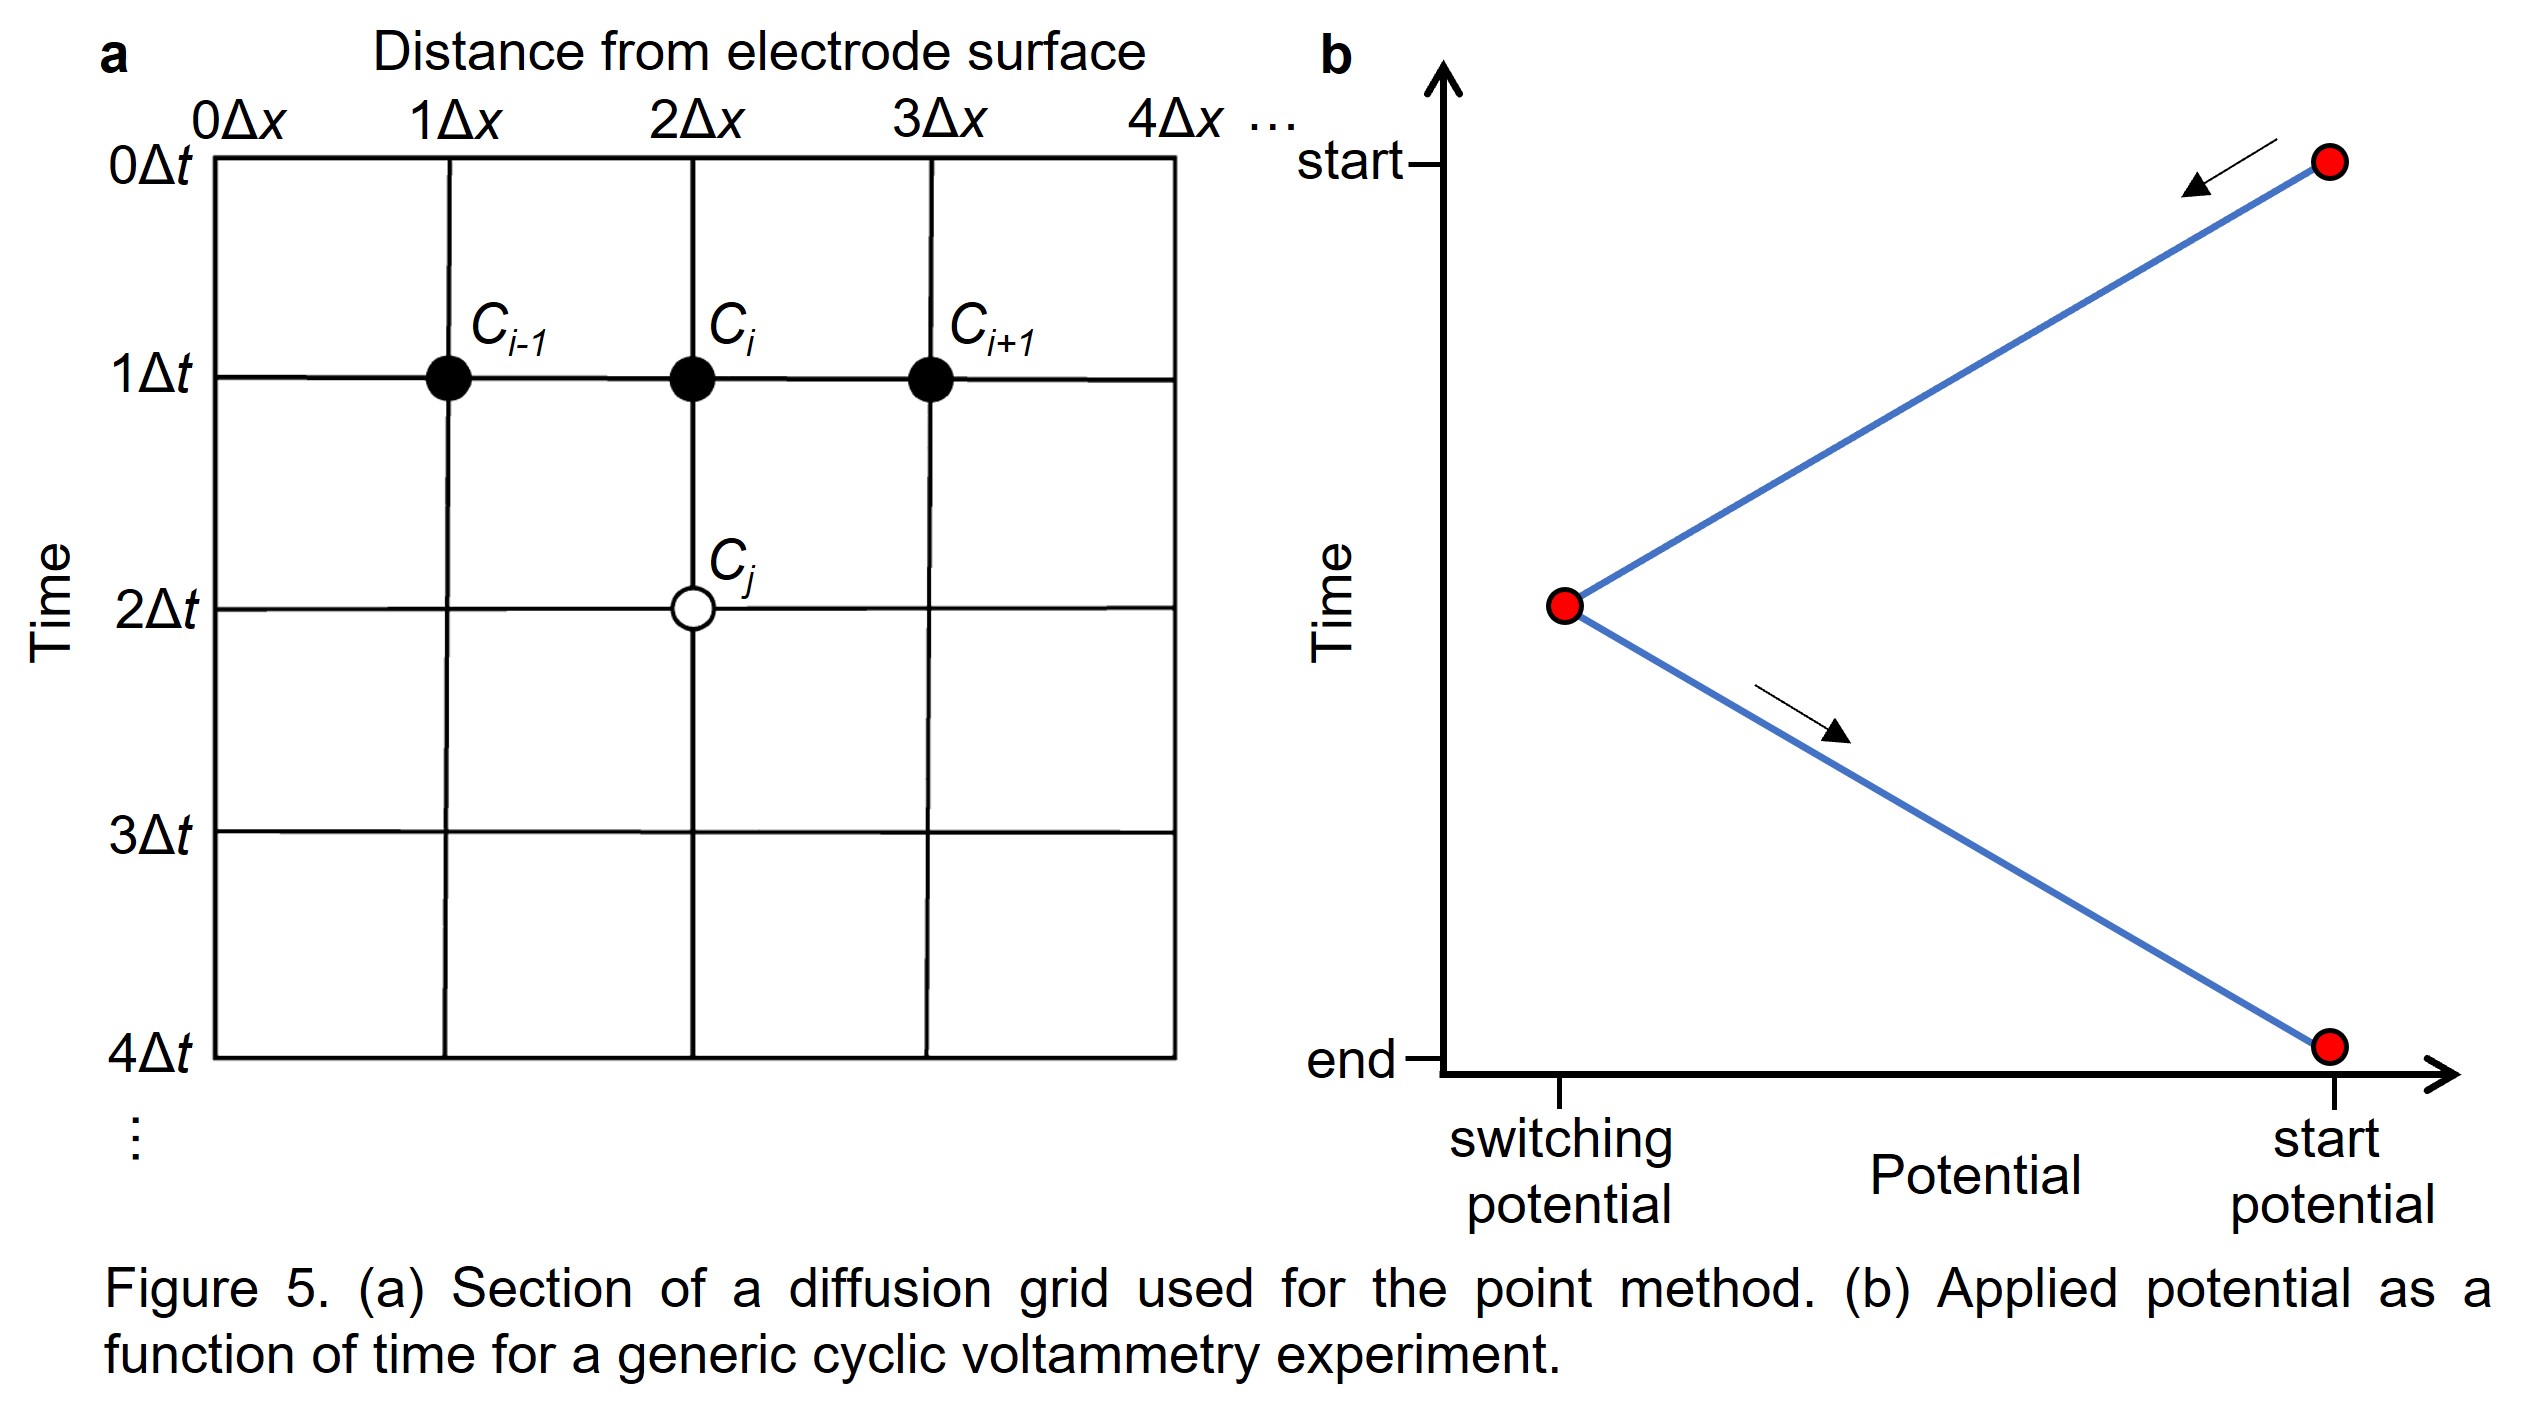
The first column represents the concentrations at the electrode surface, which are calculated values discussed below. The boundary conditions are applied in order to use the point method: 1) the last column contains concentrations furthest from the electrode surface and is thus filled with the bulk concentration; 2) the top row of the grid contains concentration values at the start of the simulation, and it is filled with the bulk concentration as well. Eq. 14 is then used to fill the rest of the grid (it should be combined with consumption or production due to redox reactions, otherwise there will be no diffusion and you will fill the grid with bulk conditions everywhere).
Results and Discussion
We are almost there, I promise. We now can work out the diffusion in its discrete form, but we need to bear in mind that the redox reactions also modify the concentration of $\mathrm{Ox}$ and $\mathrm{Red}$ at the electrode surface, and the redox current is the one my colleagues record in the CV measurement. In this sense, the redox reactions cause the concentration gradient at the electrode surface, and drive the diffusion:
\[J_\mathrm{Ox}=-D[C_\mathrm{Ox,1}-C_\mathrm{Ox,0}]/\Delta x\]where $J$ is the flux. This equation says that if the electrode consumes $\mathrm{Ox}$, $\mathrm{Ox}$ diffuses toward the electrode. Unfortunately, this equation cannot be used directly in the simulation because the concentration of $\mathrm{Ox}$ at the electrode surface $(C_\mathrm{Ox,0})$ is still unknown. However, this equation can be rearranged to $C_\mathrm{Ox,0}=C_\mathrm{Ox,1}+(J_\mathrm{Ox}\Delta x/D)$. Similarly, $C_\mathrm{Red,0}=C_\mathrm{Red,1}+(J_\mathrm{Red}\Delta x/D)$. Furthermore, the flux of the oxidized form can also be written as a rate equation $(-J_\mathrm{Ox}=k_fC_\mathrm{Ox,0}-k_bC_\mathrm{Red,0})$. Substituting the two previous equations into it gives
\[-J_\mathrm{Ox}=\frac{k_fC_\mathrm{Ox,1}-k_bC_\mathrm{Red,1}}{1+\frac{k_f\Delta x}{D}+\frac{k_b\Delta x}{D}}\] \[J_\mathrm{Ox}=-J_\mathrm{Red}\]where $k_f$ and $k_b$ are obtained from eqs. 11 and 12 at each applied potential. The total faradaic current that corresponds to each time increment is finally calculated as:
\[i_\mathrm{Total}=-nFSJ_\mathrm{Ox}\]where $S$ is the active area of the electrode. One more thing to note: aside from the diffusion we now have the equations that describe the modification of concentrations at the electrode surface:
\[C_\mathrm{Ox,0}=C_\mathrm{Ox,1}+\frac{J_\mathrm{Ox}\Delta x}{D}\] \[C_\mathrm{Red,0}=C_\mathrm{Red,1}+\frac{J_\mathrm{Red}\Delta x}{D}\]Eqs. 19 and 20, along with eq. 14, dictate the whole concentration profile from the electrode surface to the bulk solution.
The algorithm is therefore as follows: 1) use eq. 16 to calculate the flux $J_\mathrm{Ox}$, and eq. 18 to convert it to the current (the $y$ values in CV measurement), 2) use eqs. 14, 19, and 20 to update concentration profiles (after $\Delta t$), 3) repeat steps 1 and 2 until the sweep cycle is finished. A typical simulated cyclic voltammogram is shown in Figure 6, along with the concentration profiles of $\mathrm{Ox}$ and $\mathrm{Red}$. Download sample MATLAB code here.
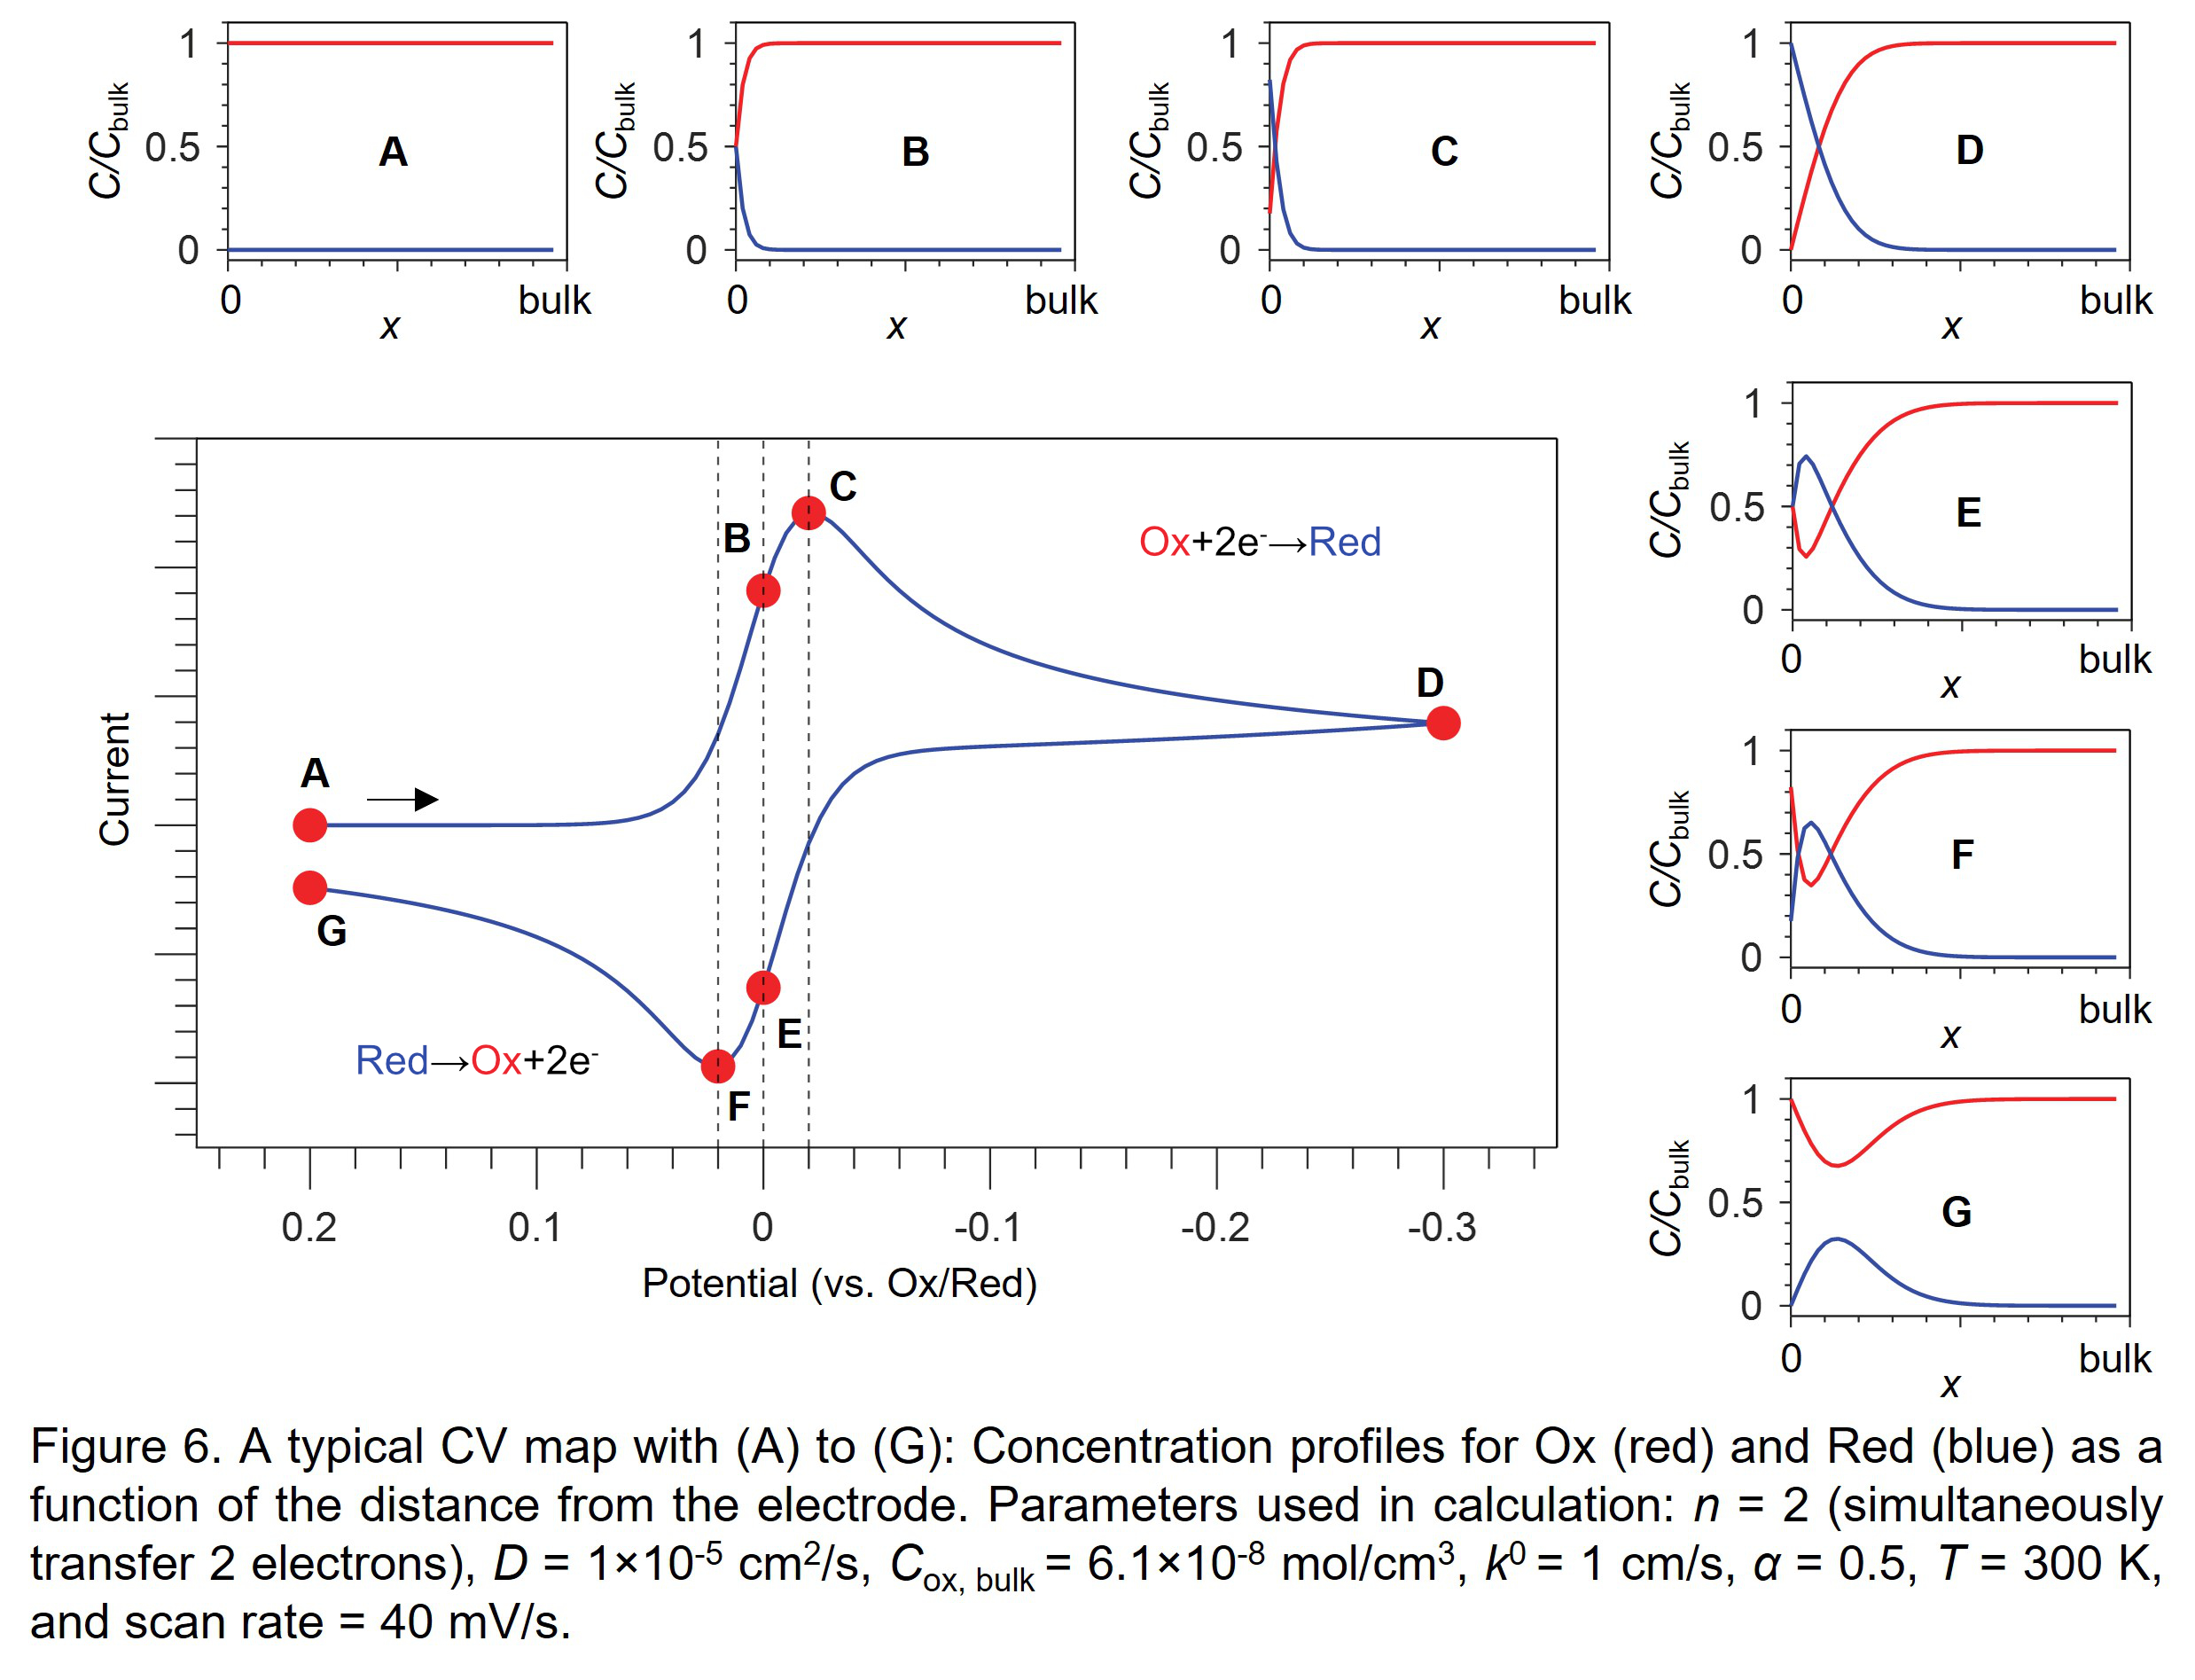
Remember what got us here: why are there two $y$ values for one $x$ value? Now we have the answer: it is not like an ordinary circuit where the potential (and resistance) dictates current. In a CV measurement, the current originates from the redox reactions at the electrode surface; its sign depends on the type of reaction (oxidation or reduction), and its magnitude depends on the reaction rate. Therefore, the potential itself is just one parameter that controls the reaction type and rate; other parameters, such as the initial condition in the EC cell and diffusion, also play important roles in determining reaction rates and thus can result in two current values for a single potential value.
References
- Elgrishi, N. et al. A practical beginner’s guide to cyclic voltammetry. J. Chem. Educ. 95, 197–206 (2018).
- Bard, A. J., Faulkner, L. R. & White, H. S. Electrochemical Methods: Fundamentals and Applications (Wiley, 2022).
- Brown, J. H. Development and use of a cyclic voltammetry simulator to introduce undergraduate students to electrochemical simulations. J. Chem. Educ. 92, 1490–1496 (2015).